Molecules on a Leash: Fast Drug Discovery express
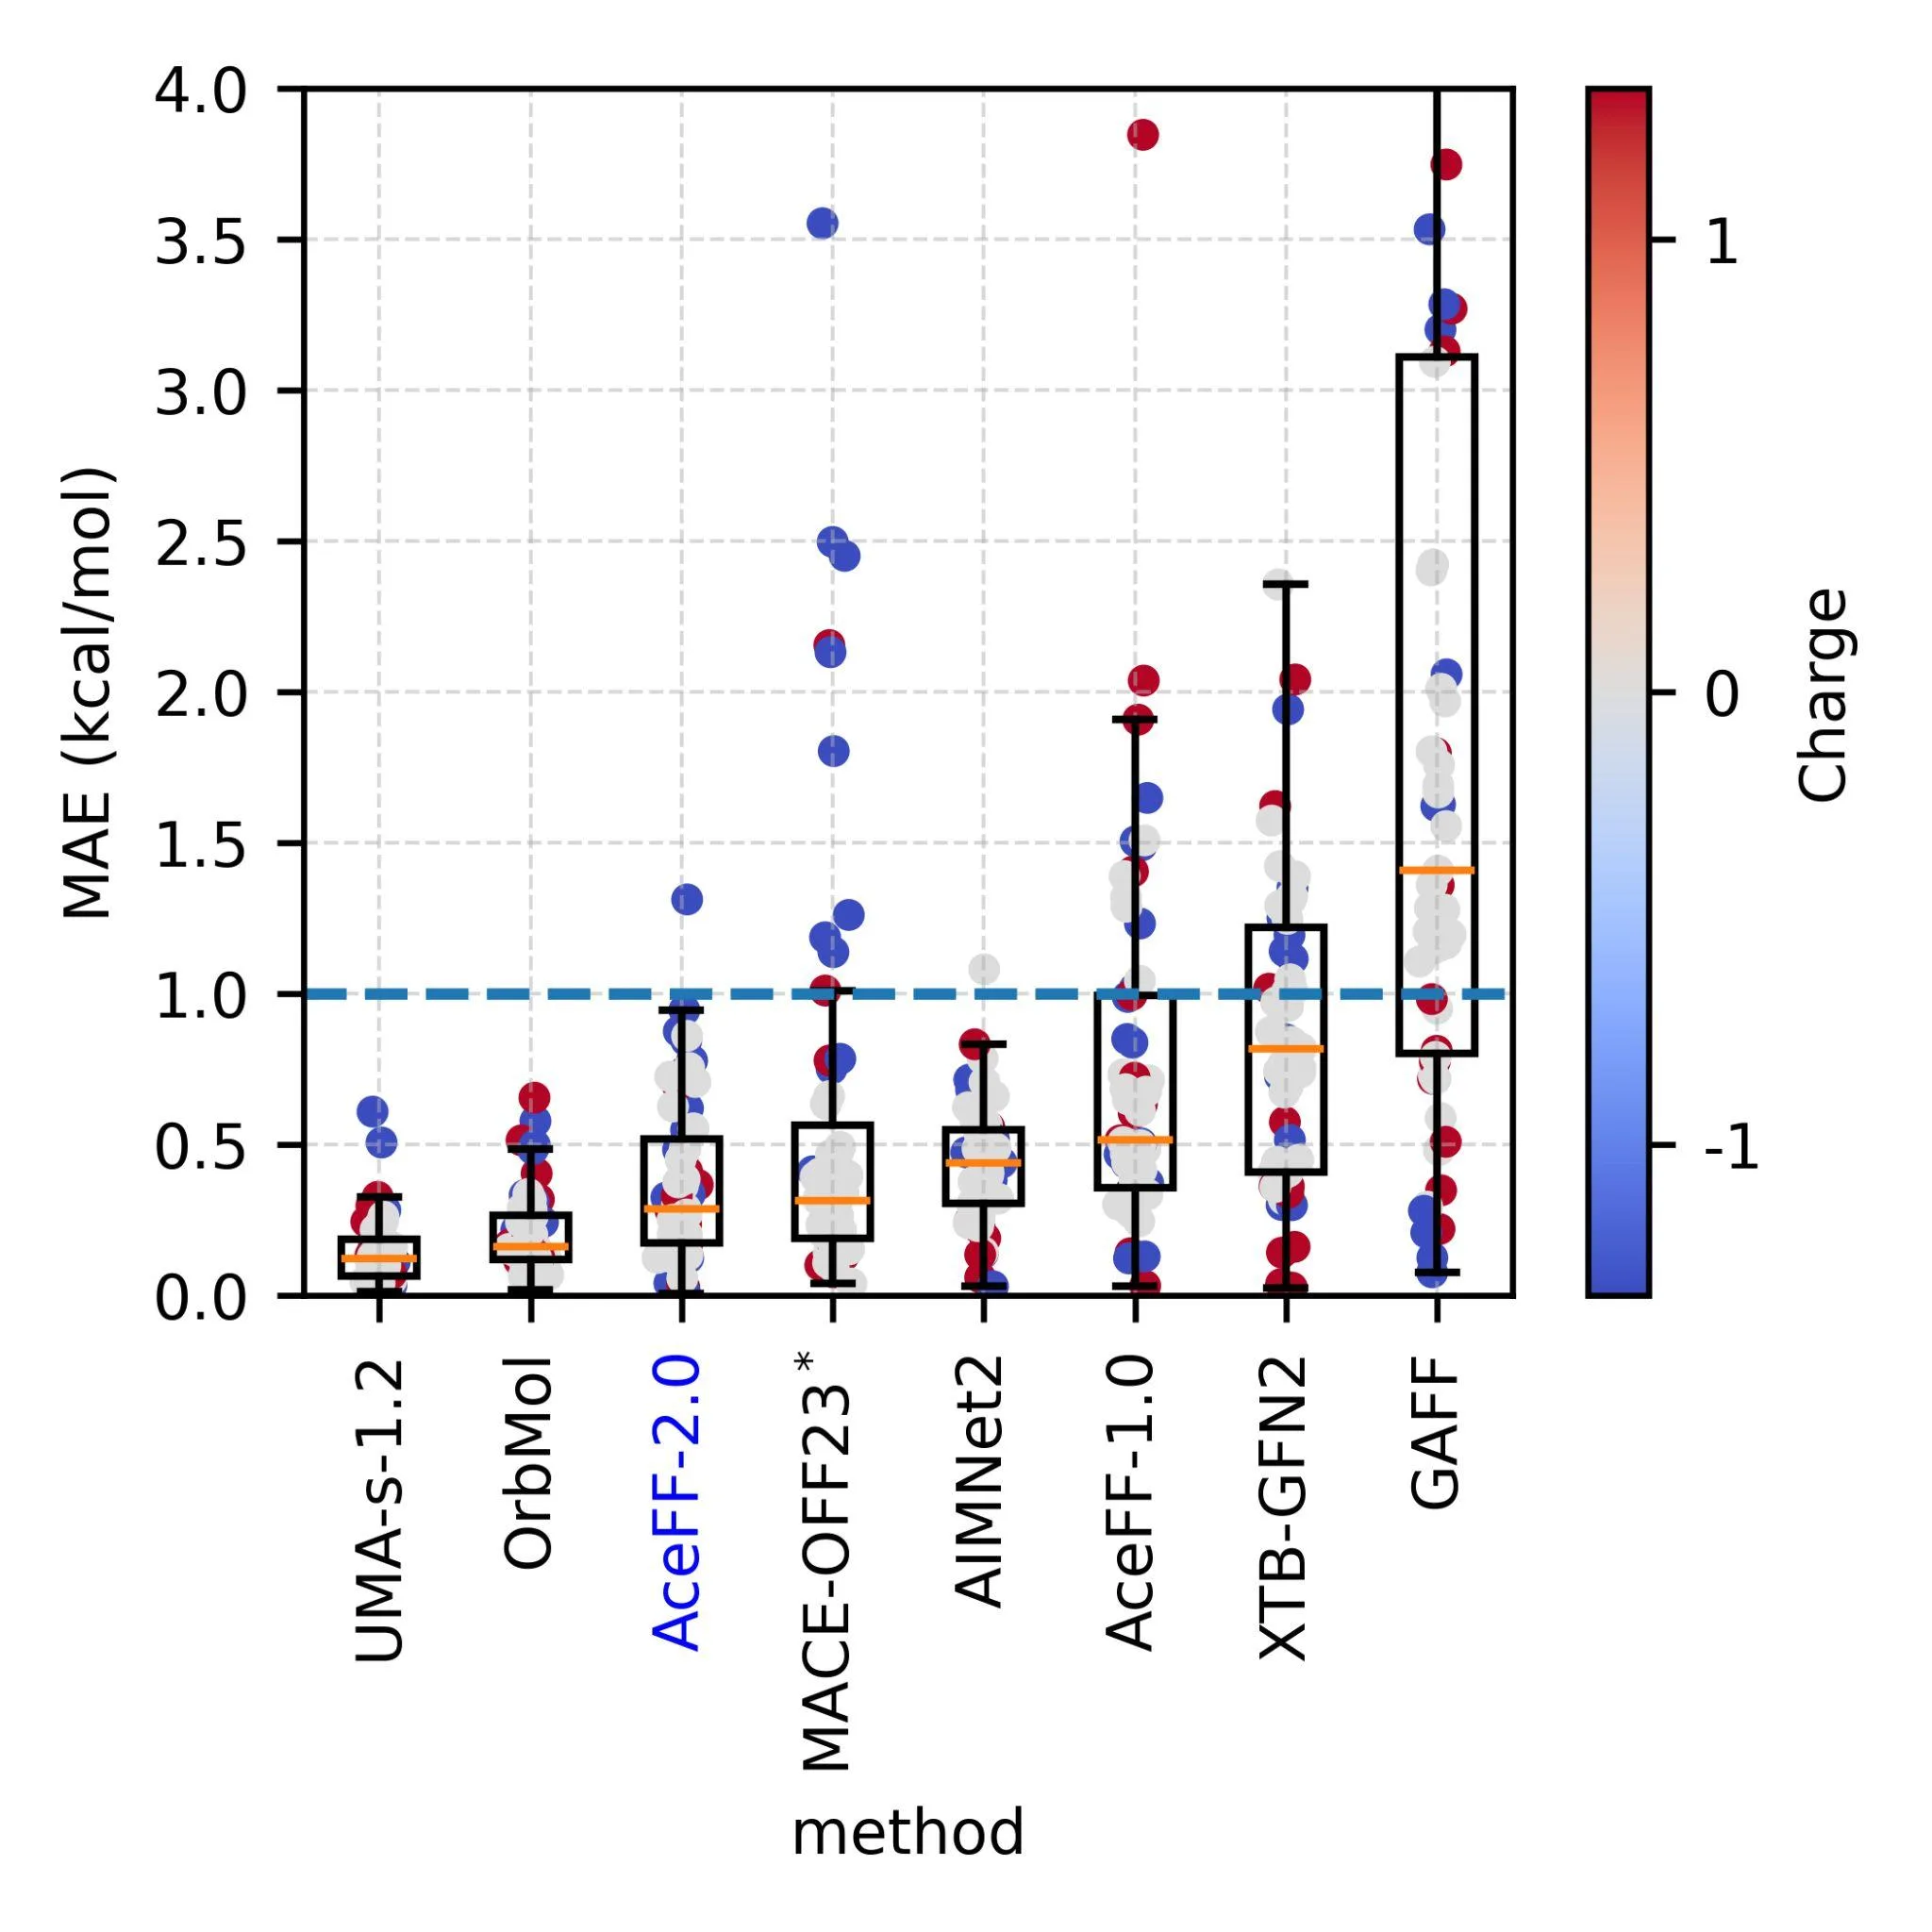
You need to sift through millions of keys to find the one that unlocks the lock of a disease. Each key is a molecule, and the lock is a protein in our body. Checking each key manually is impossible, so chemists use computer models that predict how a molecule will behave in reality. Until recently, accurate calculations took hours or days on powerful computers. The new tool AceFF does it in seconds.
AceFF is a pre-trained model on a huge dataset that understands how atoms in molecules attract, repel, and rotate around bonds. It was trained on examples of molecules similar to those used in drugs: with carbon, nitrogen, oxygen, sulfur, and other elements. The key point is that the model doesn't just memorize but derives general rules, so it even handles new, unseen molecules. In tests, it showed accuracy on par with the most rigorous quantum calculations but works thousands of times faster.
Finding a new drug is like finding a needle in a haystack. You need to test millions of candidates, and if each calculation takes an hour, the process stretches over years. AceFF reduces this time to minutes. The model is already open-source, and any researcher can download and use it in their work. It's like giving every chemist a personal supercomputer that fits in their pocket.
This approach changes the very idea of drug discovery. Instead of synthesizing and testing hundreds of compounds in the lab, scientists can first run millions of candidates through a computer, select the most promising ones, and only then move to experiments. This saves years of work and millions of dollars.
Original abstract
as written by the authors arXiv:2601.00581
We introduce AceFF, a pre-trained machine learning interatomic potential (MLIP) optimized for small molecule drug discovery. While MLIPs have emerged as efficient alternatives to Density Functional Theory (DFT), generalizability across diverse chemical spaces remains difficult. AceFF addresses this via a refined TensorNet2 architecture trained on a comprehensive dataset of drug-like compounds. This approach yields a force field that balances high-throughput inference speed with DFT-level accuracy. \mbox{AceFF} fully supports the essential medicinal chemistry elements (H, B, C, N, O, F, Si, P, S, Cl, Br, I) and is explicitly trained to handle charged states. Validation against rigorous benchmarks, including complex torsional energy scans, molecular dynamics trajectories, batched minimizations, and tests of force and energy accuracy, demonstrates that AceFF is state-of-the-art for organic molecules in the accuracy and speed regime important for drug discovery. The AceFF-2 model weights and inference code are available at https://huggingface.co/Acellera/AceFF-2.0.
The AceFF model was trained on a dataset that includes over 1.5 million molecular configurations: it's like learning the behavior of all residents of a small town to predict their movements.