Catching the Moment a Protein Breaks express
A mountain pass connects two valleys. In one, the protein is folded; in the other, it is unfolded. To get from one to the other, you must cross the saddle — the highest point on the path. In protein science, this point is called the transition state, and the coordinate that shows the distance to it is the committor. But a real protein has thousands of dimensions, the rotation angles of atoms, and finding that saddle is nearly impossible.
The authors rewrote an old variational principle so that you no longer need to impose rigid boundary conditions. Instead, they introduced a single normalization — the so-called "fidelity." Now you can use any trial function, even one that doesn't vanish at the start and equal one at the finish. This is like working with a rough sketch of the pass instead of a precise map — the key is that the scale is right.
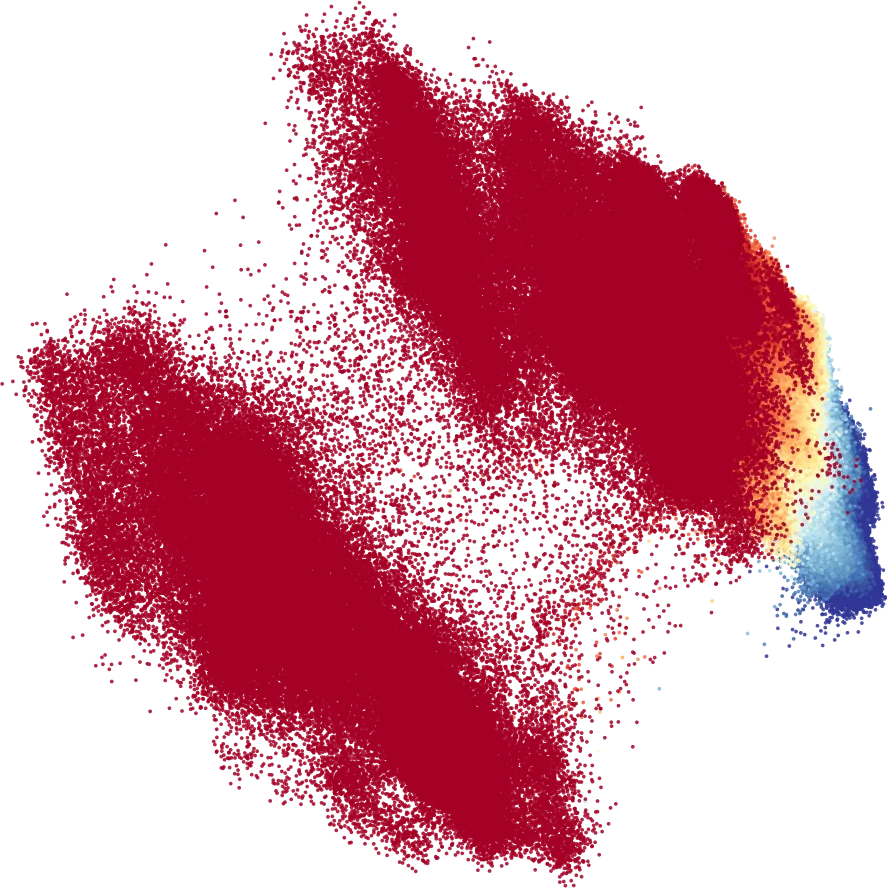
The method was tested on three proteins: AIB9, villin HP-35, and chignolin. For the first two, they obtained committors in the full space of angles, and for chignolin, folding and unfolding rates using only data from umbrella sampling. Everything computes quickly, via a closed-form formula. This opens the way to studying rare events in large systems — from protein folding to crystal formation.
The mountain pass no longer needs to be stormed blindly. A simple elevation map is enough.
Original abstract
as written by the authors arXiv:2608.02536v1
The committor is the optimal reaction coordinate for a rare transition: it pinpoints the transition state and fixes the rate, and it governs events from protein folding to crystal nucleation. It minimises a Dirichlet energy, whose value at the minimum is the reactive flux, over functions that vanish on the reactant state and equal one on the product state. In high dimensions such a trial space is very hard to build. Here we rewrite the variational principle so that the boundary conditions are replaced by a normalisation of one boundary observable, the fidelity. Any trial function is then admissible, including functions that cannot satisfy the boundary values at all. On this basis we estimate the high-dimensional committor and rates from one-dimensional profiles along projected coordinates, taking as input only pre-existing equilibrium or reweighted configurations, a diffusion constant estimate, and the two state definitions. The optimum has a closed form that is cheap to evaluate. We obtain committors for AIB9 and villin HP-35 in full torsion space, and folding and unfolding rates for chignolin from umbrella sampling alone.
Villin HP-35 folds in just a few microseconds — one of the fastest known folding processes.