How a Seven-Metal Alloy Develops a Memory express
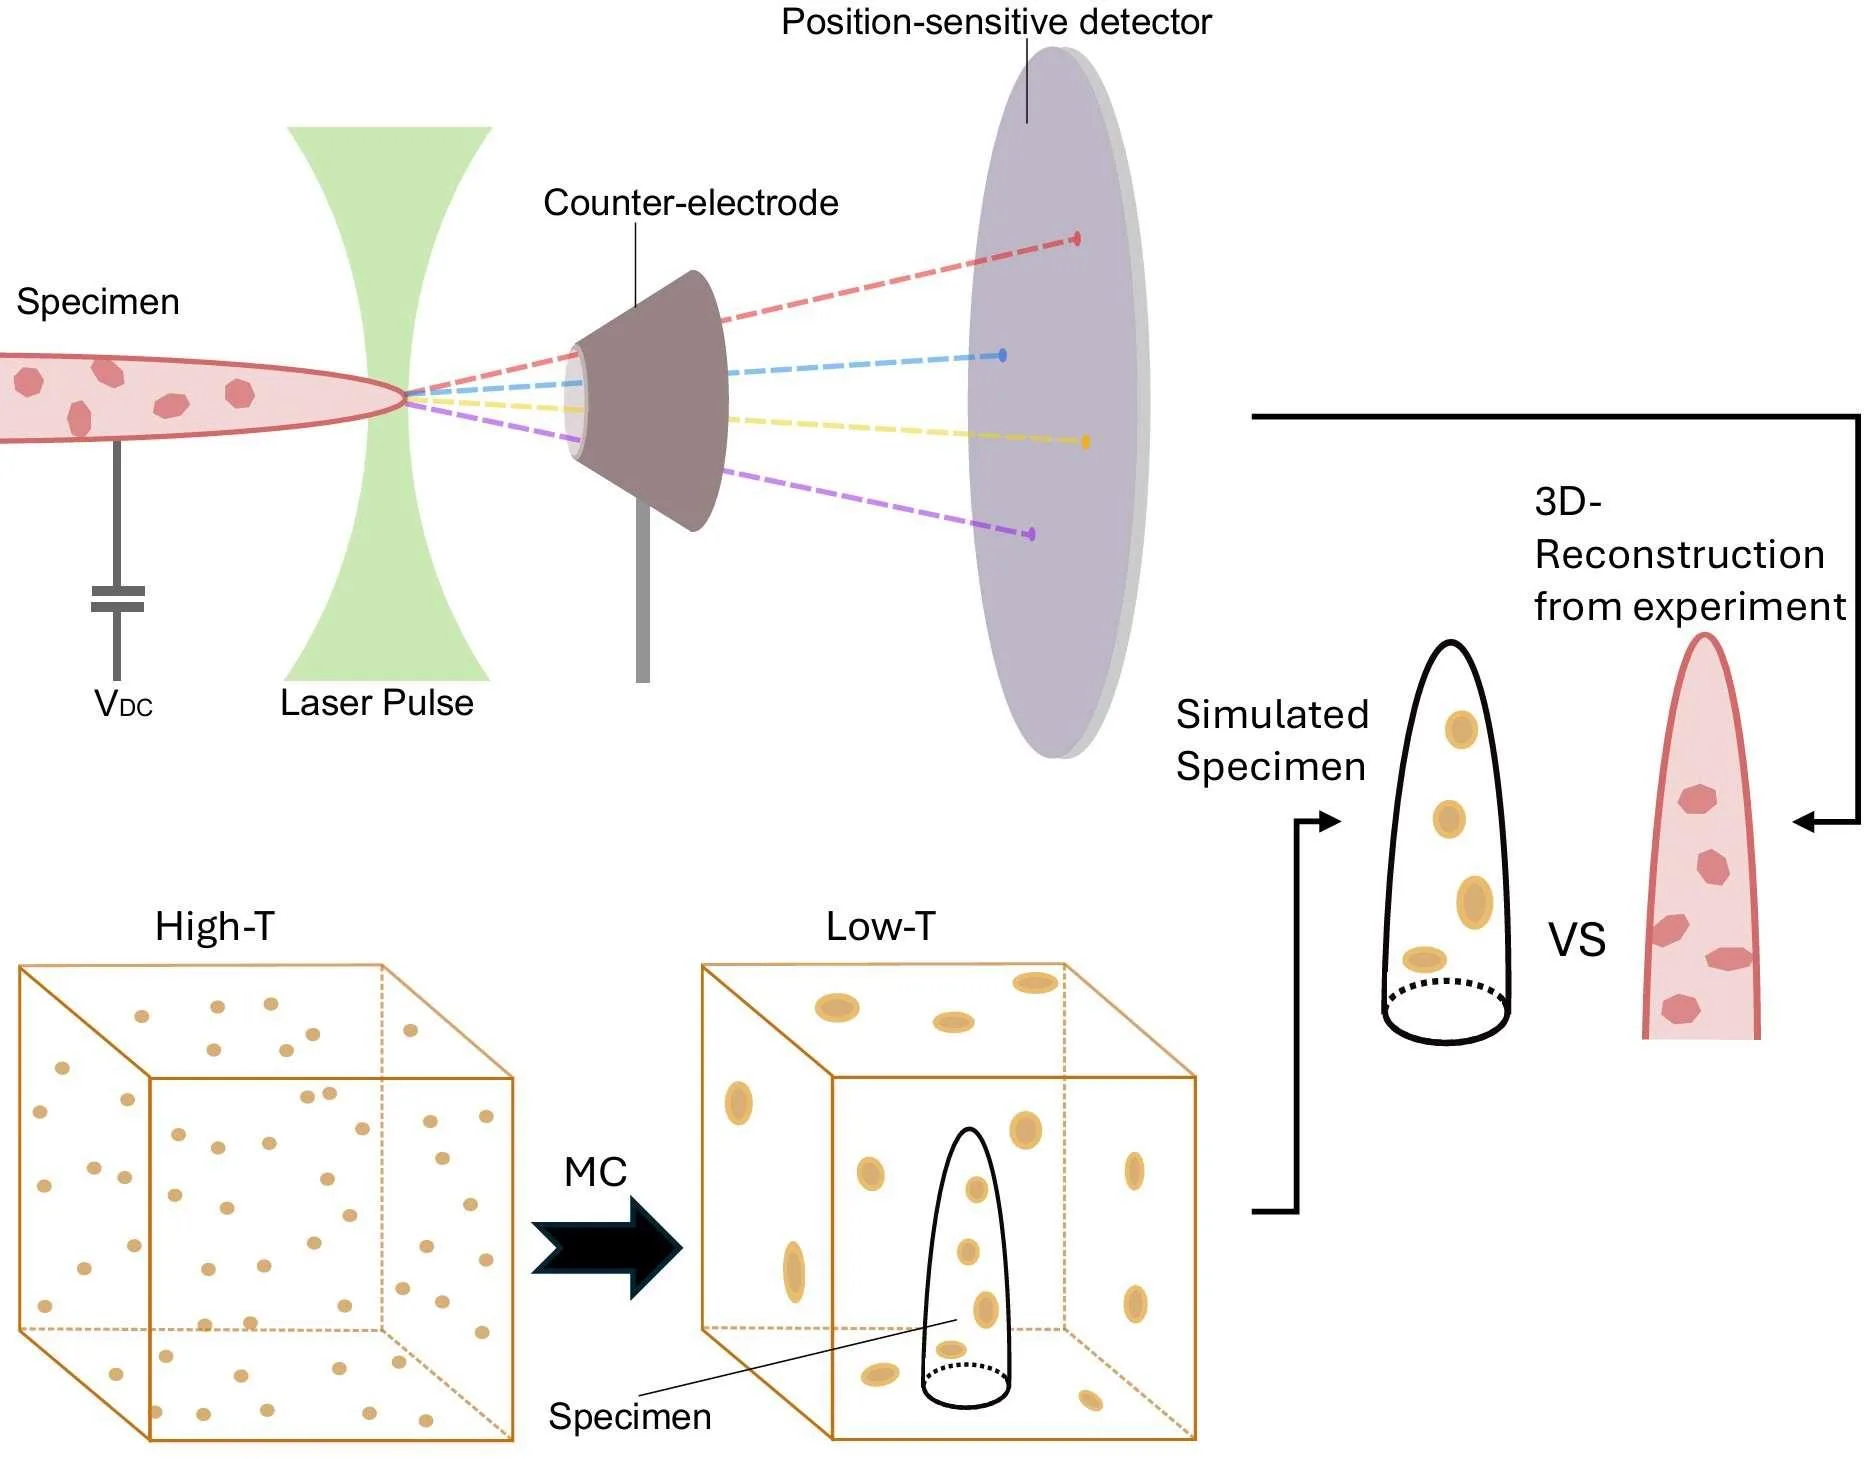
A seven-metal alloy is like a team where every player is pulling the blanket their own way. Atoms of iron, cobalt, nickel, aluminum, titanium, tantalum, and vanadium can either fall into a strict pattern or scatter into random positions. This determines whether the material will be as strong as steel or as light as aluminum. Scientists want to know in advance which pattern the atoms will choose, so they can create alloys with the desired properties.
Conventional computer calculations are too slow to track how atoms rearrange over long periods. So the authors turned to machine learning: an algorithm learns from examples and then predicts atomic behavior much faster. They compared several types of such algorithms on data from 10,000 atomic configurations. It turned out that some algorithms are more accurate, others are faster, and still others are better at capturing how atoms influence each other over distances.
The researchers also tested how important it is to account for the fact that atoms shift slightly and the lattice seems to breathe. It turned out that if you don't allow atoms to relax after packing, predictions get worse. Now materials scientists have a guide for choosing the right algorithm for modeling high-entropy alloys. This brings us closer to creating materials that will serve in jet engines or nuclear reactors, where conventional alloys quickly degrade.
Original abstract
as written by the authors arXiv:2603.21207
Tailoring the performance of next-generation high entropy materials requires a deep understanding of the competition between entropy-driven random solid solution and enthalpy-driven chemical ordering. Investigating such order and disorder complexity demands atomistic simulations that achieve high accuracy, efficiency, and generalizability across vast spatial, temporal, and especially chemical scales. While machine learning (ML) interatomic potentials have transformed molecular dynamics, they remain limited in capturing diffusion-driven chemical evolution over long timescales. The recently introduced SMC-X method brings exciting opportunities. Realizing its full potential requires a comprehensive study, which is the focus of this work. To assess model performance, we systematically benchmark invariant and equivariant architectures using a density functional theory dataset of more than 10,000 configurations spanning seven elements: Fe, Co, Ni, Al, Ti, Ta, and V. To understand the roles of pairwise and higher-order interactions, we decouple their contributions across chemical space using an explainable machine learning approach. We also examine the impact of lattice relaxation by comparing models trained on datasets with and without structural relaxation. Our results clarify how to choose ML surrogate models for Monte Carlo simulations, bridge the gap between theory and experiment, and lay a foundation for establishing ML-accelerated Monte Carlo as a computational microscope for chemical complexity.
High-entropy alloys are sometimes called 'cocktail alloys': you mix five or more metals in nearly equal proportions, and you get a material with unexpected properties — for example, simultaneously strong and ductile.