How to Measure a Molecule's Energy More Precisely Than the Instrument Allows express
A quantum computer measures a molecule's energy much like an old tape measure that only has markings every meter. If the actual length of a table is 2.37 meters, the tape shows “two and a bit,” and you can't do better. Physicists call this resolution: the step of the grid, finer than which you cannot go.
The authors of the new work turn this flaw into an advantage. Instead of a single measurement, they make several, each time shifting the start of the ruler slightly — by a centimeter, two, five. Then they combine all the coarse readings and average them. The effect is reminiscent of the old trick of shifting the scale: each individual reading is imprecise, but together they allow recovering the true length with an error much smaller than the scale division.
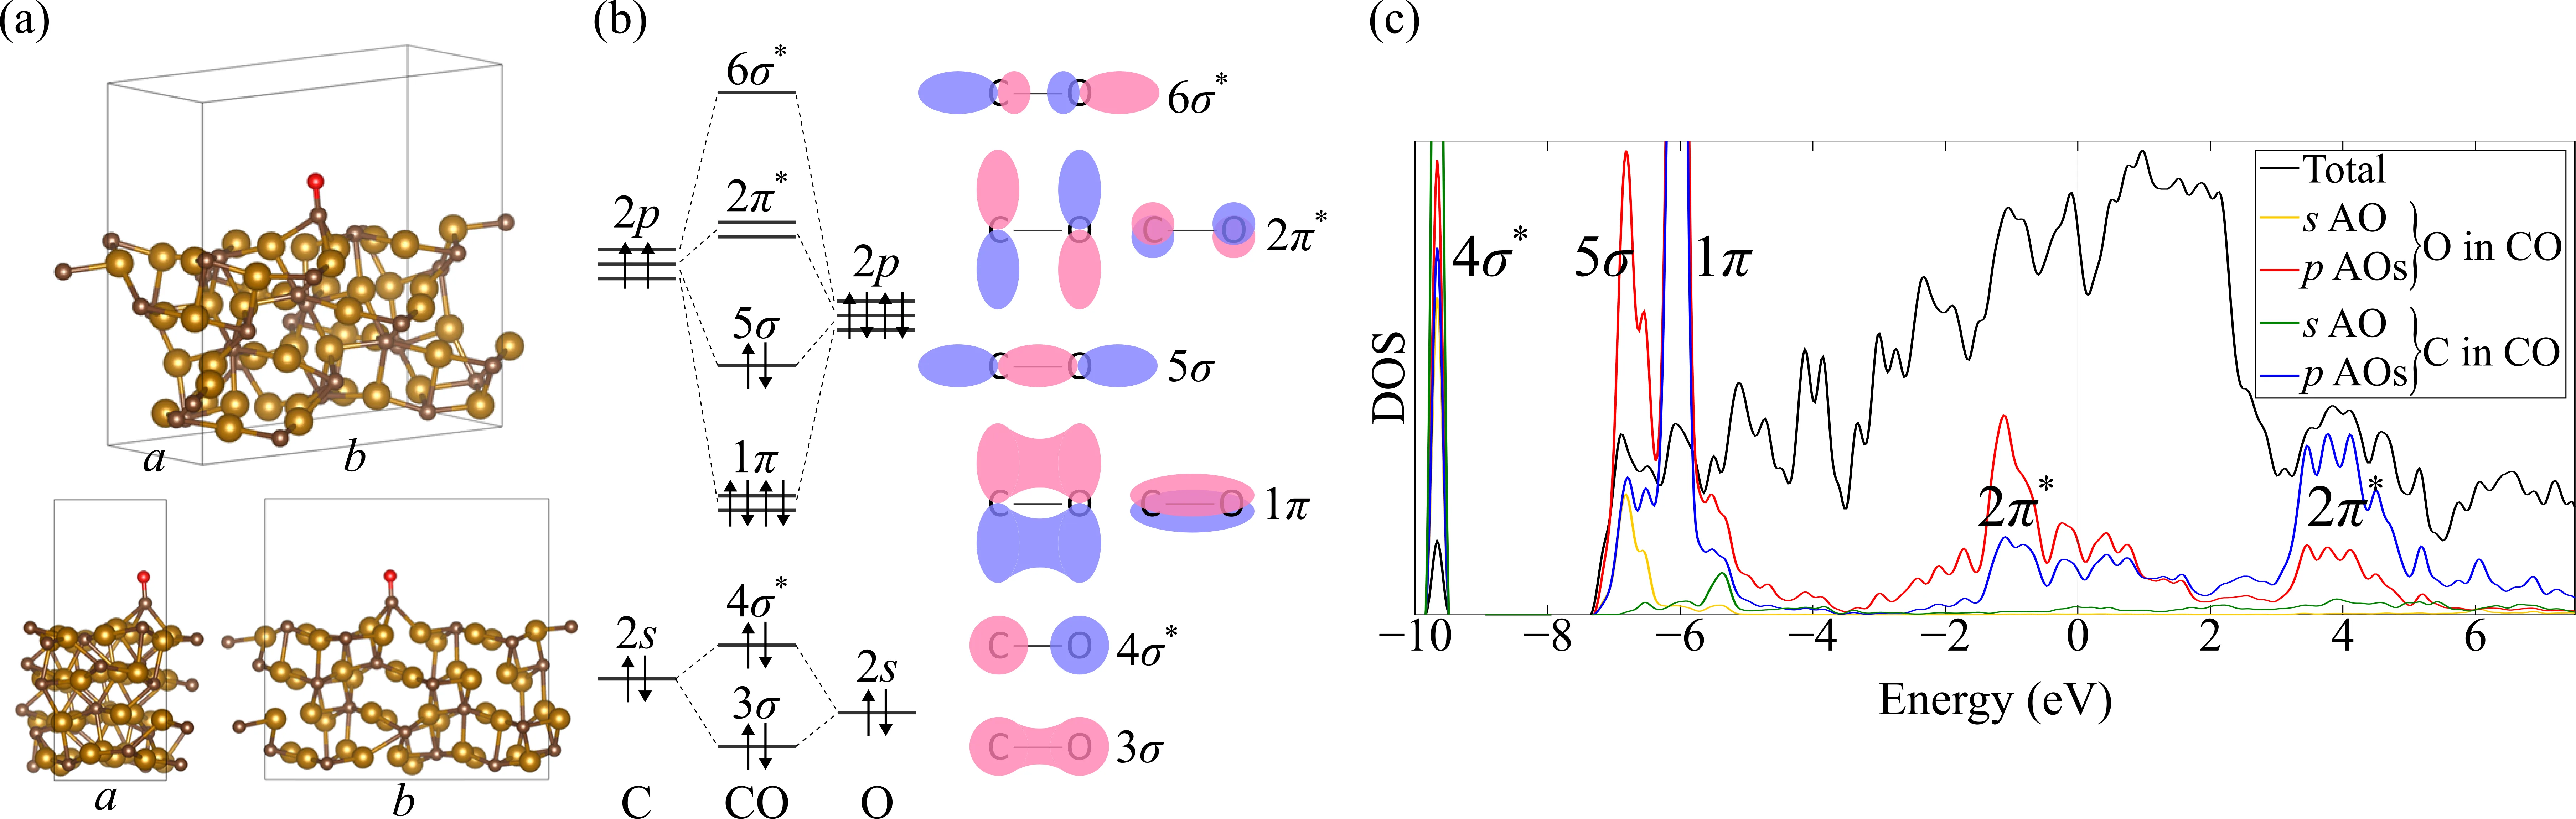
The method was tested on a real problem: a carbon monoxide (CO) molecule sticks to the surface of an iron catalyst, like a magnet to a fridge, but it's held not by magnetism but by electron clouds. Such systems are important for the chemical industry, where catalysts speed up reactions, but calculating their behavior on a regular computer is very hard. A quantum computer should do better, but for now it's noisy and error-prone.
The researchers ran their scheme on the Quantinuum H2-2 quantum processor in two variants: on physical qubits (which are prone to errors) and on logical qubits protected by Steane's error-correcting code. Even when the measurement histograms were heavily noisy, the “shifted ruler” method reliably recovered the molecule's energy spectrum. Moreover, averaging over shifted grids smooths out the “ravines” in the mathematical landscape of the search, preventing the algorithm from getting stuck in a false minimum, like driving on a bumpy road, but the suspension smooths out the potholes, allowing you to keep the right direction.
This approach paves the way for calculating complex chemical systems on quantum computers that haven't yet achieved perfect accuracy. Instead of waiting for flawless hardware, scientists are learning to squeeze the most out of what exists today.
Original abstract
as written by the authors arXiv:2605.29674
Quantum phase estimation (QPE) is an underlying technology for extracting the excitation spectra of many-electron systems, yet its practical use on current hardware is hindered by low grid resolution and environmental noises. Here we propose QPE averaged over variable grids (QAVG), a vernier-type approach that combines low-resolution QPE with multiple origin shifts and physically motivated continuous parametrization to reconstruct the spectra accurately. We introduce this approach into an end-to-end workflow for the {\it ab initio}-based model system for a CO molecule adsorbed onto the $$\chi$$-Fe$$_5$$C$$_2$$ surface. We perform experiments on Quantinuum H2-2 using both physical QPE circuits and logical QPE circuits encoded in the Steane code with offline bit-flip correction. We demonstrate that QAVG accurately reconstructs the spectra with deviations much smaller than the nominal QPE resolution, even when the noisy histograms are used. The cost landscapes averaged over the shifted grids substantially suppress the local minima arising from the spectral leakage, thereby stabilizing the optimization of trial parameters. These results indicate that QAVG provides a robust route to quantum simulations of correlated spectra toward the era of early-fault-tolerant quantum computers.
The vernier (or nonius) was invented in the 17th century for astronomical instruments, and now this principle helps quantum computers measure molecular energies more precisely.