Neural Network Learns to Read Molecules by Their Carbon Signature express
Every carbon atom in a molecule carries a passport — the energy with which its deepest electrons sit in their orbitals. This energy is sensitive to neighbors: add an oxygen nearby or a double bond, and the number in the passport changes. Chemists have long measured such energies with X-rays, but calculating each new molecule takes hours of machine time.
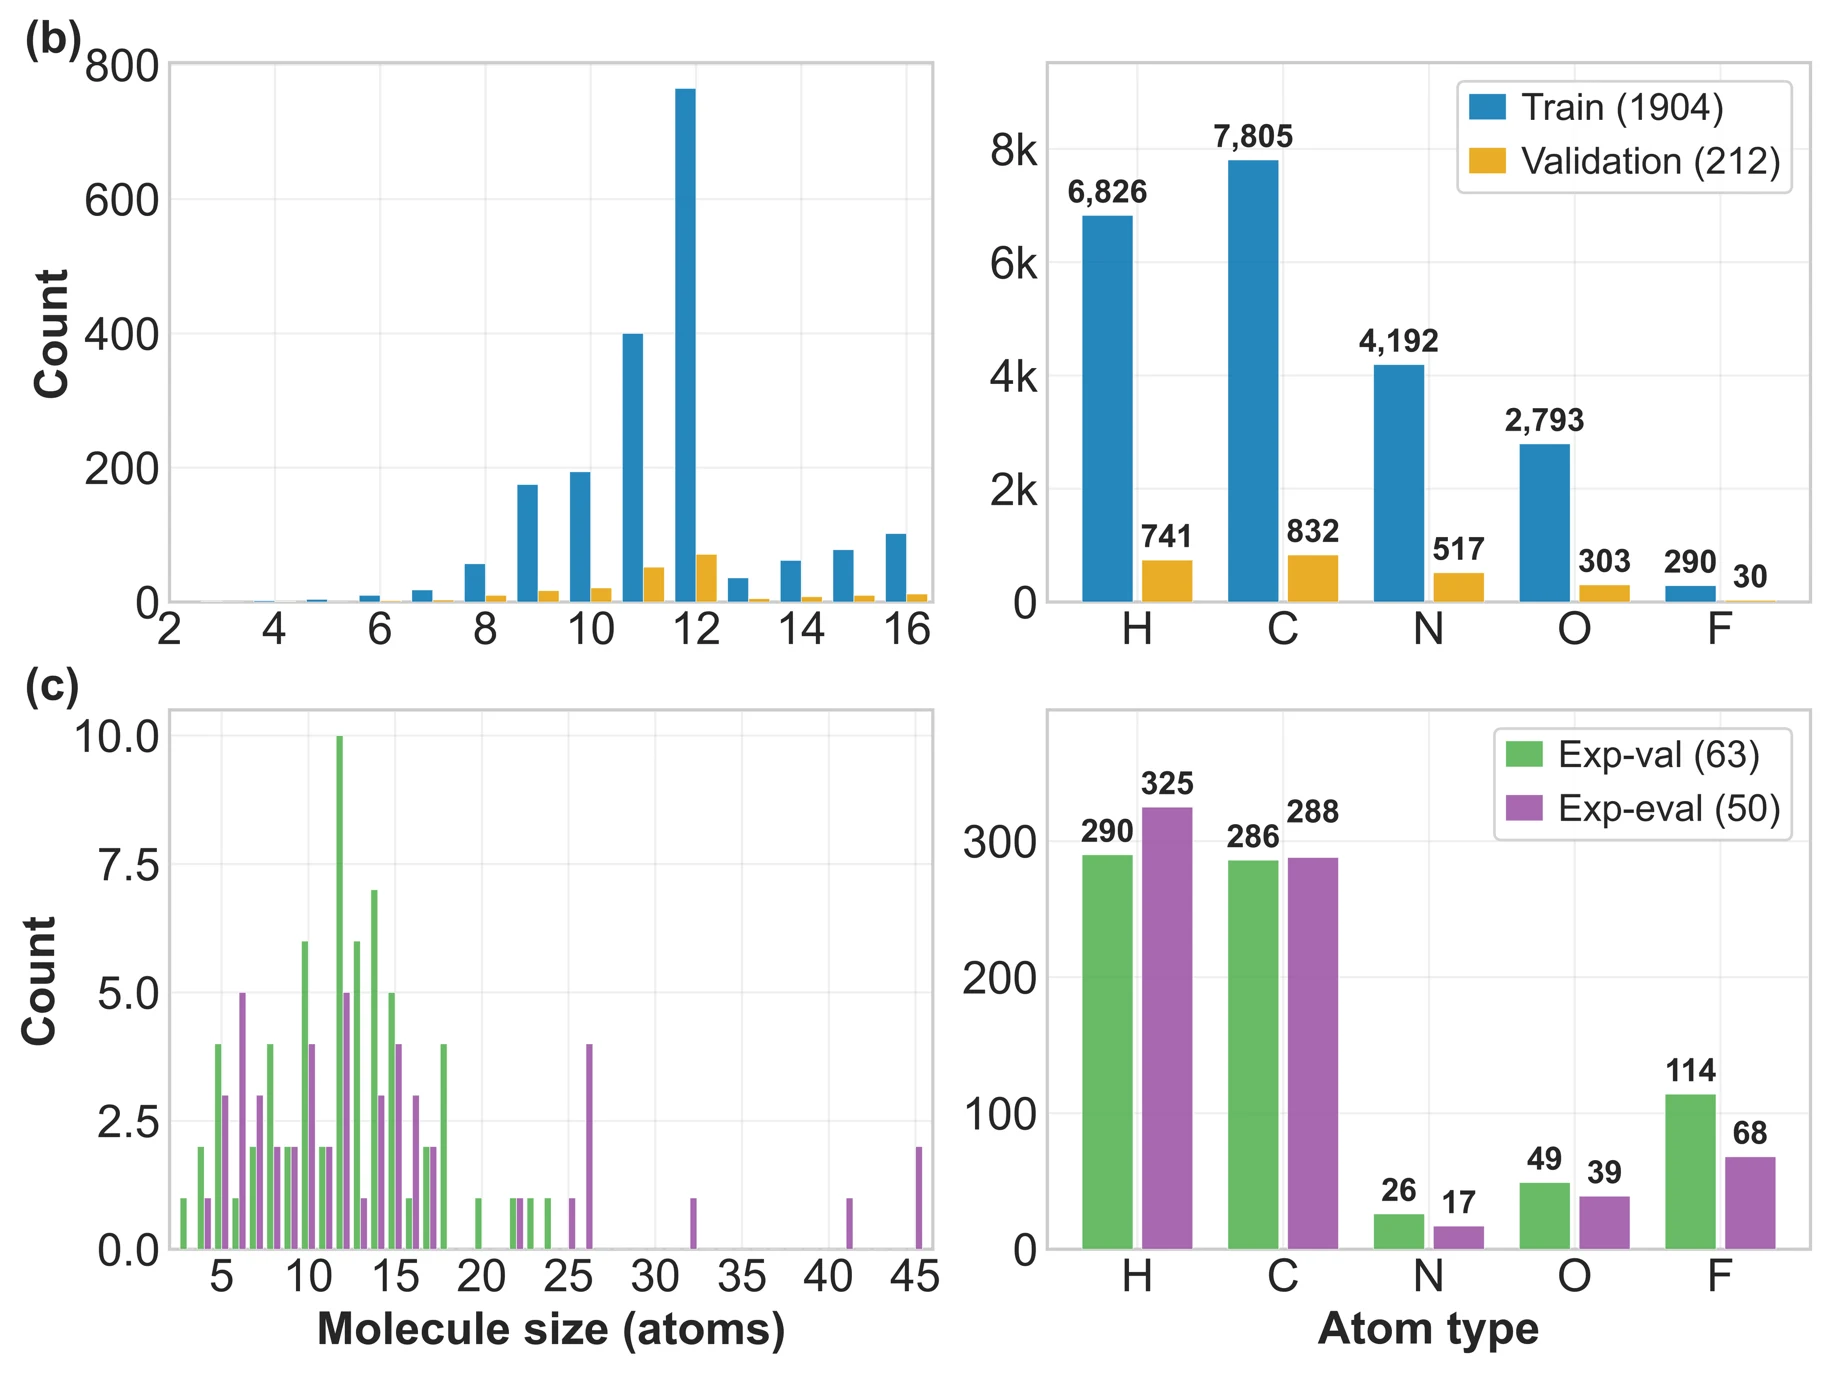
The new model works like a seasoned archivist who, from a single photo of a building, instantly determines the strength of its foundation. It sees a molecule not as a jumble of balls but as a graph — a network where atoms are nodes and bonds are edges. The neural network was trained on 8637 carbon centers in 2116 molecules ranging from 4 to 16 atoms, using data from multiconfigurational density functional theory (a sophisticated quantum calculation method). When tested on 570 real measurements in 113 molecules, some reaching 45 atoms, the average error was 0.33 electronvolts. That's roughly one hundredth of the energy of visible light.
A special feature of the model is chemically meaningful node features that reduce dependence on the depth of message passing in the graph. In simpler terms, the neural network doesn't need to 'ask' distant neighbors many times to understand an atom's environment. It's like gauging a person's mood not from their entire chain of conversations but from a few key words.
The model is also robust to distorted geometries: those that arise from thermal vibrations or early stages of a chemical reaction. It was tested on tautomers of avobenzone (45 atoms)—molecules that shift a hydrogen atom and are linked to UV protection. The neural network instantly distinguished the forms and produced energies consistent with experiment. This paves the way for rapid screening of drugs and materials.
And here's the curious thing: carbon's passport changes by several electronvolts depending on neighbors — enough to distinguish graphite from diamond by X-ray spectrum alone.
Original abstract
as written by the authors arXiv:2604.27070
Graph neural networks are promising architectures for fast, accurate and transferable predictions of core-electron binding energies, which depend on the local bond environment. Here we present a graph neural network model for predicting carbon 1s core-electron binding energies in organic molecules. The model is trained with multiconfiguration pair-density functional theory on 8637 carbon atoms in 2116 molecules with 4-16 atoms and evaluated against 570 experimental values in 113 different molecules containing 3-45 atoms. Previous work benchmarked a mean absolute error of 0.27 eV to experiment for the training data level of theory [J. Phys. Chem. A 2025, 129, 36, 8419-8431] and the present model demonstrates an experimental evaluation error of 0.33 eV with good size transferability to larger organic molecules. An equivariant graph neural network is benchmarked against its rotationally invariant analogue and a model comprised of the smooth overlap of atomic positions descriptors and kernel ridge regression for training data efficiency and stability to non-equilibrium geometries absent from the training data. All models show good training data efficiency and the graph based models have improved transferability to non-equilibrium geometries. The use of chemically informed, graph-normalized node features reduces the graph neural network's dependence on message passing depth. A case study on the 45 atom avobenzone tautomers demonstrates the model's ability for instant and precise analysis of complex molecules. The software and data are provided by the open-source AugerNet package at https://doi.org/10.5281/zenodo.19689244.
The energy of carbon's core electrons changes by several electronvolts depending on neighbors — enough to distinguish graphite from diamond by X-ray spectrum alone.